Table of Contents
Quick answer
Capture the report promptly, preserve product and lot identity, record current stock status and identify the responsible escalation route. Keep observations separate from conclusions. Reporting obligations depend on the event, market and the organization’s role; a supplier investigation should not delay a required report.
A customer reports an unreadable label, an unexpected product characteristic or a suspected device problem. The first message may be incomplete. Waiting for a perfect description can lose time, while immediately attributing a cause can distort the record.
The useful first response is to capture what is known, preserve identity and send the concern through the appropriate quality and regulatory route. This article concerns the product record and distribution response, not clinical management of an affected patient.
What should be captured at intake?
Record the reporter’s description in neutral language, including when the issue occurred and when the business became aware of it. Keep observed facts separate from assumptions about manufacturing, handling or use.
A practical intake form includes the product reference, lot or serial information where available, quantity involved, photographs or relevant documents, location of the product and a contact for follow-up. If a field is unknown, mark it unknown rather than guessing from a previous order.
Collect only personal information needed for the legitimate follow-up and handle it through the appropriate controlled channel. Avoid circulating patient details in a general sales thread. A commercial account contact and a product lot may be sufficient for the first operational escalation.
How does medical device complaint traceability connect to inventory?
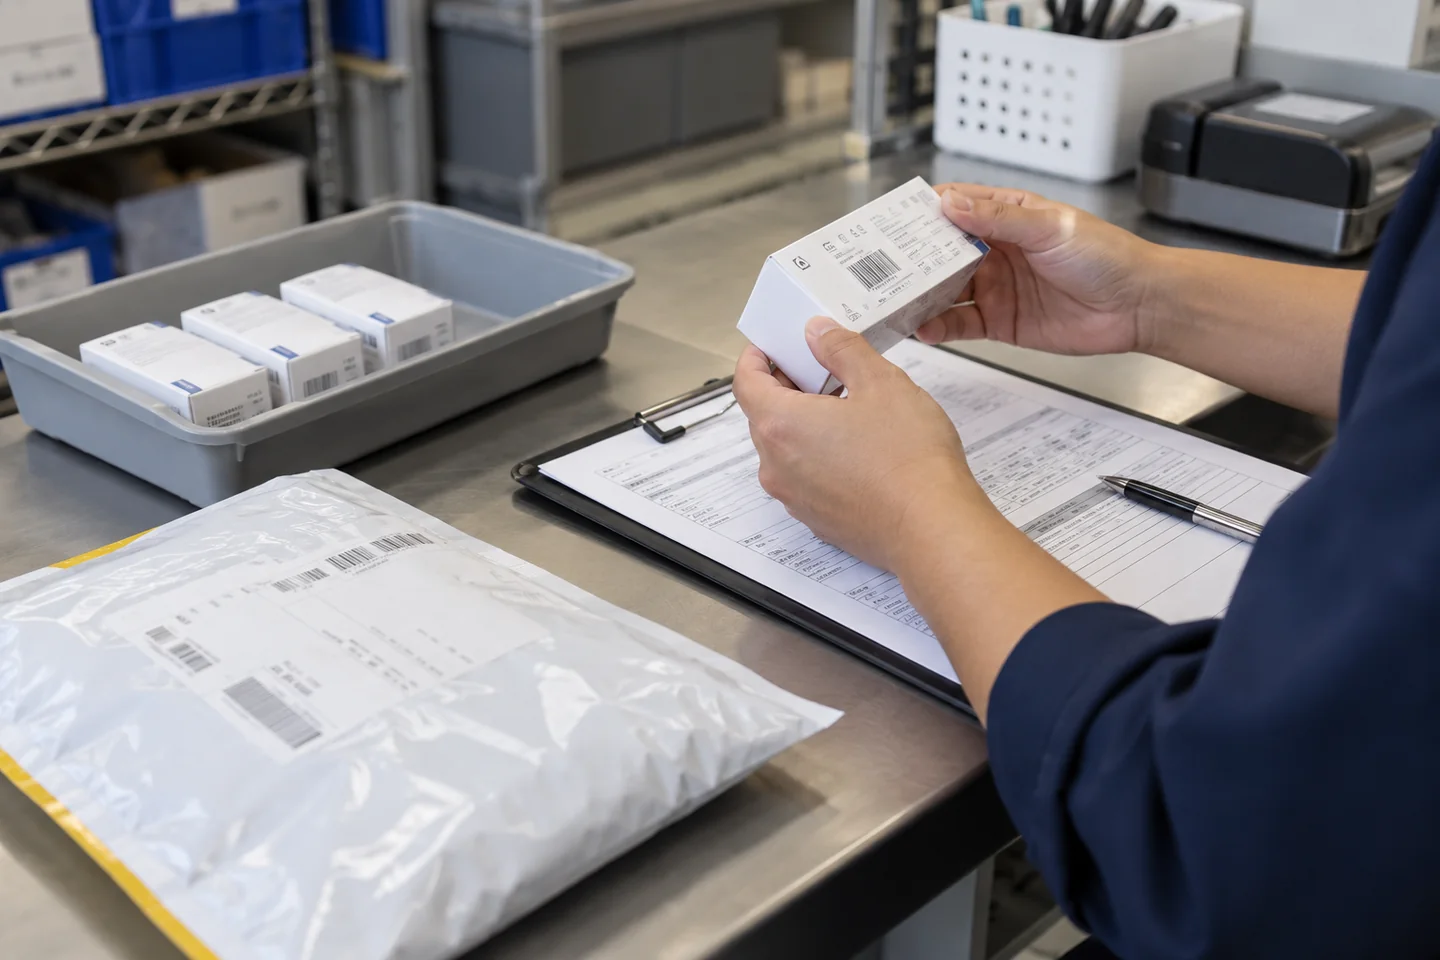
Link the intake record to receipts, remaining stock and dispatches for the identified product and lot. The objective is to establish scope, not to assume every unit is affected.
Consider a hypothetical reconciliation for a lot of 600 units:
| Recorded status | Units | Evidence to retrieve |
|---|---|---|
| Held at distributor | 180 | Location and hold record |
| Sent to Customer A | 240 | Dispatch record and recipient |
| Sent to Customer B | 160 | Dispatch record and recipient |
| Retained evaluation units | 20 | Internal allocation record |
| Total accounted for | 600 | Reconciliation against receipt |
These invented quantities illustrate an accounting check only. If the records account for 590 rather than 600, investigate the ten-unit difference; do not insert an unexplained adjustment to make the table balance. Preserve transfers, returns and other movements that may explain the gap.
The UDI and lot-record guide explains the underlying data structure. The complaint workflow tests whether those records can be retrieved when the business needs them.
When does regulatory reporting enter the workflow?
Early enough to meet the applicable obligation. Reporting depends on the nature of the event, the market and the organization’s role. FDA’s mandatory medical-device reporting overview distinguishes requirements for manufacturers, importers and user facilities. A distributor should not assume that one reporting rule applies to every participant or every country.
Assign the assessment to the responsible regulatory function and document the route used. A commercial response from a supplier does not, by itself, discharge another party’s duties. An investigation can continue while required notifications proceed; do not make a completed root-cause report an automatic prerequisite for escalation.
What should happen to stock and evidence during review?
Apply the appropriate hold and handling controls to stock within the defined scope. Retain relevant packaging, labels and records, and follow safe instructions for any physical return. Do not alter the complained-about unit in an attempt to reproduce a problem outside an authorized investigation.
Record communications with dates, recipients and requests. If the supplier expands the scope to additional lots, update the inventory search and recipient list accordingly. Keep superseded findings distinguishable from the current assessment.
FDA’s recalls, corrections and removals overview explains a separate set of concepts and obligations for the US. A complaint and a recall are not interchangeable labels. The responsible parties must determine the appropriate action from the evidence and applicable requirements.
How can a distributor test readiness without a real incident?
Run an internal retrieval exercise using a historical lot and clearly mark it as a simulation. Ask the team to identify the remaining stock, recipients, supporting documents and escalation contacts. Do not send simulated recall notices to customers.
Document missing fields and slow handoffs, then correct the workflow. The result should be a process that can reconstruct the distribution history without relying on one salesperson’s memory. Keep the complaints channel and the agreed supplier contacts accessible so an incoming concern reaches the right function promptly.
Frequently Asked Questions
Should a complaint wait until the customer sends the physical product back?
No. Record and escalate the available information promptly, then supplement it as further evidence arrives. Physical return arrangements should follow the relevant safety and quality procedures and should not delay required reporting.
Does every complaint mean that a recall is needed?
No. A complaint is an input to assessment and investigation. Any corrective action or recall decision must consider the facts, product risk and applicable requirements; neither automatic dismissal nor an automatic recall is an adequate assessment.